
Proving that an answer is the answer
Before this page
Thermochemistry §4, where the minimization and its multipliers are set up, and Standard states.
An interior-point method minimizes
For a consistent Gibbs potential, the formulation being certified is the one set out in Thermochemistry §4: minimize
Which equilibrium is computed
A system is said to be at equilibrium when none of its properties changes with time and when, after a small disturbance that is later removed, it returns to the same state (Anderson and Crerar, 1993) (§3.3). The definition is operational, and it is met by states that are not the minimum of the Gibbs energy: aragonite kept at room temperature, or a mixture of hydrogen and oxygen, persists on the observation timescale although calcite and water lie lower. Such a state is metastable, separated from the stable one by an energy barrier that the conditions do not allow the system to cross, whereas the stable equilibrium is the lowest state compatible with the constraints.
A third kind of state is neither. A metastable phase such as aragonite is in internal equilibrium: its temperature, pressure and composition fix its properties, and only its transformation into another phase is blocked. A glass, or a crystal whose distribution of cations over its sites stopped changing on cooling, is not: its configuration froze when it could no longer follow the temperature, its properties depend on that history, and one more variable is needed to describe it (Richet, 2001; Secs. 2.4a and 6.3a, pp. 39 and 143–144). Properties that do not change with time are therefore no proof of equilibrium, and a minimization over amounts has no variable for such a state; the glass of a slag enters a calculation as a reactant of prescribed extent (The enthalpy of a glass).
A minimization knows nothing about barriers, and the state it returns is the stable equilibrium of the system that was posed. Metastability therefore enters a calculation only through the way the system is posed, and it does so in two ways. A species absent from the list cannot form, so that leaving it out computes the equilibrium that is metastable with respect to it. The first calculation of Getting started leaves out dissolved hydrogen, oxygen and methane for this reason: with them, the minimization would also settle the oxidation state of carbon, that is a redox equilibrium between carbonate and methane which water at room temperature does not reach without a biological catalyst, and which the example has no intention of describing. Choosing the species list treats the same decision for cement phases, where a missing phase is a far more common error than a deliberate exclusion. The second way is to withdraw some amounts from the budget altogether, which is what the coupling with kinetics does with the phases it integrates in time.
The latter is a partial equilibrium, in which only a subset of the possible transformations has reached its balance while the others proceed at their own pace. It is the whole principle of the coupling between kinetics and equilibrium (Kinetics under partial equilibrium), where the aqueous speciation and the precipitation of hydrates are taken as instantaneous while the dissolution of the clinker phases follows rate laws. Each instant of such a trajectory is then the stable equilibrium of a smaller system, the one left once the kinetic amounts have been withdrawn from the budget, and it is this equilibrium that speciated_states recomputes and certifies. The phase rule says what such a system may hold. At temperature and pressure imposed, the phases coexisting at a stable equilibrium are generically no more numerous than the independent components, the rank of the conservation matrix; every transformation held back by kinetics adds one independent component (Richet, 2001; Secs. 9.1a–b, pp. 197–199), which is why the unreacted clinker can stand beside its hydrates. A last distinction concerns space. A system out of equilibrium as a whole may be made of regions each at equilibrium, which is called local equilibrium; every calculation of this package is zero-dimensional and describes one such region, transport between regions being outside its scope (The water budget of a hydrating paste argues what this excludes for a paste).
The certificate below checks stationarity, conservation and phase conditions for the posed system. These prove a stable global equilibrium when the model defines a convex Gibbs potential, as qualified below. Whether the species list, fixed amounts and boundary conditions describe the experiment is a separate modeling question.
Why the question has an answer
Write the Gibbs energy in RT units as G(n) = Σᵢ nᵢ μᵢ(n). Its ideal part
has Hessian
by Cauchy–Schwarz applied to G is convex, the feasible set
Optimality can then be checked: a composition satisfying the KKT conditions is globally optimal. Convexity alone does not imply a unique composition: the ideal Hessian has a scaling null direction, and pure-phase terms are linear. Uniqueness requires strict convexity on the feasible directions or additional conditions excluding degeneracy. Different starting points in a convex problem can reveal incomplete convergence or degenerate global minima; they cannot produce distinct isolated local minima of different energies.
What changes with a non-ideal model
Two requirements must be checked separately. First, the supplied potentials must be the gradient of one CapillaryWater likewise lies outside the ideal proof.
These mathematical issues differ from kinetic metastability. An activation barrier is a feature of a molecular or nucleation pathway, not a second minimum that must appear in the bulk composition objective (Metastable does not mean a local minimum of G draws the two side by side). A thermodynamic solve does not model barrier crossing. Suppressing a phase or holding kinetic amounts fixed changes the feasible set, and a constrained minimum can be globally optimal on that set while remaining metastable relative to an excluded transformation. Waiting longer changes the constraints through kinetics; it does not ask the equilibrium optimizer to climb an activation barrier.
The certificate
optimality_certificate checks the conditions below, on any composition and whatever produced it: stationarity, conservation, and a phase condition for each kind of absent or vanishing species. Writing
| condition | on which species | meaning |
|---|---|---|
interior (n > floor) | stationarity | |
| — | conservation of matter | |
| a pure phase at its bound | that phase undersaturated | |
| a member of a present phase below the floor, whose potential the interior determines | it holds no less than its potentials give it | |
| a mixing phase held entirely absent | that solution cannot form |
The last row is Michelsen's tangent-plane measure, and it is a separate test because a mixing phase needs one: its members are never exactly zero while it exists, so they are neither interior nor at a bound, and a solid solution left out of the assemblage used to pass the certificate unexamined. The trial composition is refined against the phase's own activity model, so the test is not the ideal approximation.
Two subtleties decide whether the check is meaningful.
A species at its bound obeys the inequality, not the equality. Imposing the equality on an amount held at 1e-16 whose mass-action value is e⁻³⁰⁰ misstates its log-activity by 263 RT units, and the check then reports a residual of 74 for a composition solved to 5e-12. A member of a present phase below the floor is read the same way, as the truncation of a smaller exact amount: it may hold more than that amount, not less. Excluded from every test, such a member left behind by the search would pass, as H⁺ at 3e-100 mol would in a cement paste whose potentials give it 1.2e-16.
A species carrying a vanished component is absent by the constraint, not by thermodynamics, and its saturation index is meaningless — the element potential of a component the budget does not supply is determined by nothing. The test for that is not bₖ ≈ 0 but bₖ ≈ 0 with the non-zero entries of row k sharing a sign: only then does H⁺ row carries +1 for H⁺ and −1 for OH⁻, so its zero total is the ordinary state of pure water; treating it as degenerate kills the entire acid–base system and returns pH 7.000 with the calcite undissolved.
Here n_absent_component. Measured on a site family of
What the certificate proves, and when
These conditions are sufficient for a global minimum only when the chemical potentials are the gradient of one Gibbs energy and that energy is convex over the feasible set. The first property can be measured at the audited composition, since the second derivatives of one energy commute: the Jacobian of the log activities has to be symmetric. It is for the ideal dilute model, whose solvent row is the partner of the solutes'
The second property is not a property of the point: a Hessian positive at the answer proves a minimum there and nothing elsewhere. A global scope therefore rests on a convexity established for every composition, phase by phase, the Gibbs energy being the sum of the energies of the phases. A pure phase contributes linearly; an ideal mixture (a gas, an ideal solid solution, ideal site mixing, a constant capacitance, whose charging energy is a convex quadratic of the charge) a convex function; a non-ideal solid solution what mixing_convexity decides; and the aqueous phase what follows.
A convexity bound for the Debye-Hückel form
Let every ion carry
The ideal part has the quadratic form
which is positive. The excess part
a term of rank one, negative wherever
The Hessian is therefore positive as soon as
proves the aqueous phase convex whatever its composition. At 25 °C, with
Where convexity is not proved
The certificate then asks what the answer is. The Hessian of reduced_curvature. Positive, the answer is a strict local minimum on its active set, which a non-convex model may hold beside others. Negative, a direction that conserves matter lowers the energy and the answer is a saddle: a Pitzer set whose
The certificate reports which case applies as scope, with the reasons in scope_reasons: :global_minimum when both properties hold and the constraint is variational; :local_minimum when the energy exists, its convexity is not proved and the reduced curvature is positive; :kkt_point when that curvature is not positive, or when the constraint is not variational (a retention law that shifts the activity of water, a temperature or a pressure made an unknown); and :self_consistent when the activities are not a gradient. optimal keeps its meaning in all four, the conditions being met to tolerance.
Mass action or minimization
Programs computing a speciation fall into two families, according to the data they take and the unknowns they solve for (Anderson and Crerar, 1993) (§19.2). The first writes, for every species outside a chosen basis, the law of mass action of its formation reaction with a tabulated equilibrium constant, completes these laws with the mass balances and the charge balance, and solves the resulting nonlinear system by a Newton method on the activities of the basis species; the phases allowed to precipitate are declared, and whether each of them is present is decided from its saturation index. PHREEQC (Parkhurst and Appelo, 2013) is built on this formulation. The second minimizes the Gibbs energy of the whole system under the mass balances, with the standard potentials of all species as the only thermodynamic data, so that no reaction is written and the assemblage is part of the result rather than of the input; it is the formulation of GEM-Selektor (Kulik et al., 2013), of Reaktoro (Leal et al., 2017) and of this package.
From the same data, both families compute the same state whenever both converge, which follows from the stationarity conditions of Thermochemistry §4. Taking as basis the primary species, whose potentials are the negated multipliers
which is the law of mass action of the reaction forming
The certifying solver
DualEquilibriumSolver solves the KKT system directly, in element potentials. From
Two levels. The inner one inverts the solutes' mass-action laws at fixed potentials and fixed solvent amount; the outer is a Newton on m element potentials, and the amounts of the active pure phases
A solid solution is a mixing phase without a solvent. At given potentials its composition is explicit,
each end-member taking the larger share the cheaper it is against the potentials of its elements, and the phase is present exactly when
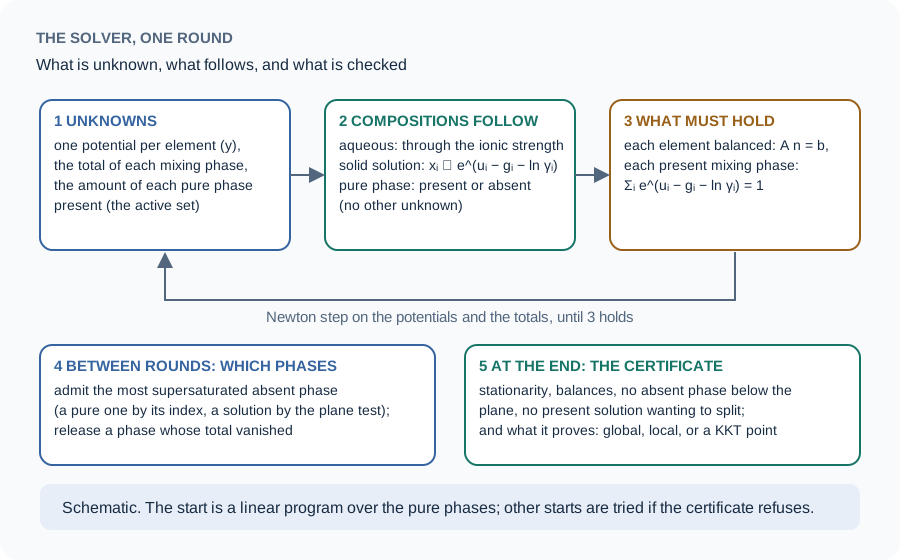
Absent phases are admitted one at a time between the rounds, the most supersaturated first, a pure phase on its saturation index and a mixing phase on its tangent-plane distance: admitting several at once feeds a cycle in which a phase is admitted, driven negative, released and admitted again. The search starts from the linear program obtained by setting every activity to one, solved over the pure phases, which also proves an impossible budget to be impossible; when the certificate refuses the answer, other starts follow, which equilibrate_certified lists.
For the Debye–Hückel, Davies and Truesdell–Jones models the inner level is one equation. Their activity coefficients depend on the composition through the ionic strength alone, so at a given
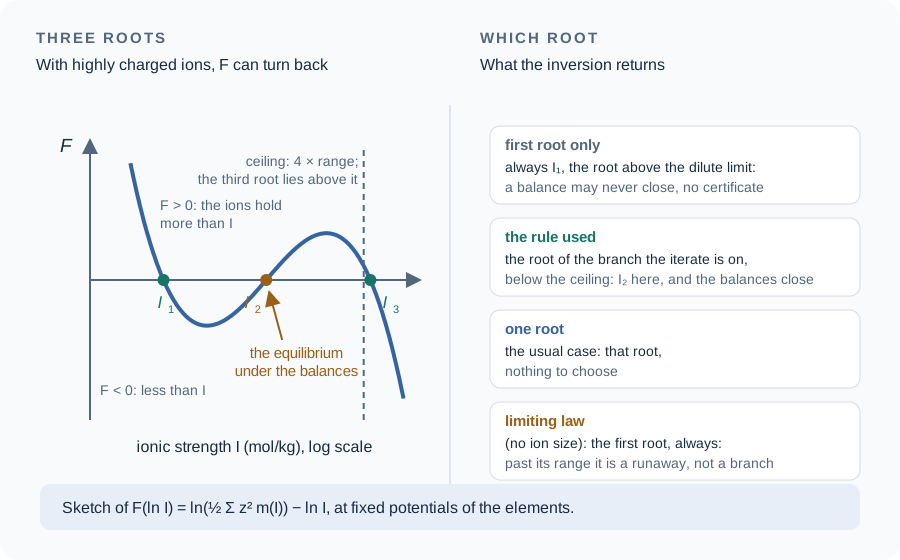
SIT and Pitzer add terms in the molalities themselves, the
The solvent is deliberately not inverted through its own mass-action law: its activity cannot exceed one, being a mole fraction in some models and y can demand more, for which no finite composition exists. It belongs to the outer system, where the balance determines it.
What it buys, measured
On calcite in pure water the certified pH is 9.90, where the package's interior-point solve alone stops at 6.96, a point its return code does not distinguish from the minimum. On the Reaktoro reference (calcite, CO₂ and water) both routes now agree with Reaktoro on every species: above 10⁻⁵ mol to 10⁻³ relative, the trace ions to 5 %, the worst being CaOH⁺ at ×1.032 on 1.6 nmol. That reference used to carry a @test_broken for CaOH⁺ at ×2.47; what closed it was the convergence test moving to the true KKT error at μ = 0 (OptimaSolver 0.4.1), and test/equilibrium_reference.jl is now 26 plain assertions.
speciated_states certifies every instant it replays and names any it cannot. On a full ordinary Portland cement over 28 days, all forty replayed instants are certified, with element balances between 1e-11 and 1e-13 mol.
Where to go next
The chapter continues with Activity models, the first of the places where a mixture stops being ideal and the one on which every aqueous equilibrium depends. The solver described here is driven in practice from the tutorial Chemical Equilibrium, and the API entries are equilibrate_certified, optimality_certificate, DualEquilibriumSolver and saturation_indices.