
Solid solutions
Before this page
Thermochemistry and Standard states §2, where the standard state of an end-member is set.
A pure solid has
This matters in cement more than anywhere else. C-S-H is not a compound with a formula but a phase of variable Ca/Si; the AFm phases exchange sulfate, carbonate and hydroxide on the same interlayer site; alkalis partition into hydrates rather than staying in solution. A model that only knows pure phases predicts sharp appearances and disappearances that are not observed.
1. Ideal mixing: the activity is the mole fraction
Put
which is entropy alone — the
That is IdealSolidSolutionModel, and it is the default. Note what it already buys: an end-member at
The curve, its tangent, and the plane of the rest of the system
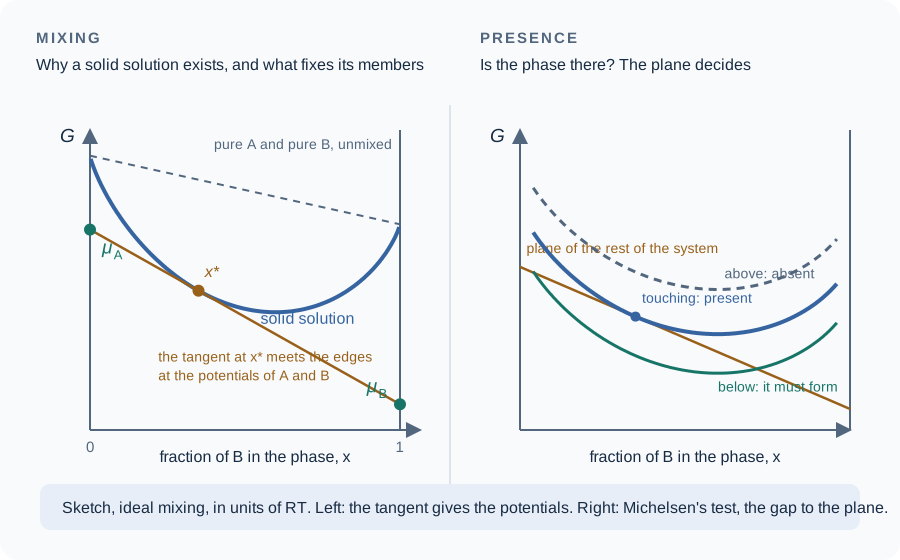
Per mole of phase, the Gibbs energy of a binary of end-members A and B is the straight line of the two end-members unmixed,
At equilibrium each element has one potential, shared by every phase, and those potentials give each end-member a potential
2. Non-ideal mixing: the excess Gibbs energy
Real end-members interact. Everything beyond the ideal term is collected into the excess Gibbs energy
and then _ss_log_activities! writes log(x[k] + ϵ) + _excess_ln_gamma(model, k, x, T) into the log-activity vector, so the two terms are visibly separate and a model supplies only the second. The model of section 7 is written differently, and has its own method.
3. RegularSolutionModel — one interaction energy per pair
The simplest non-ideal form keeps only pairwise contacts, with one energy per pair of end-members:
which is the symmetric multicomponent Margules expression, and it is _excess_ln_gamma(::RegularSolutionModel, …) line for line. For a binary it collapses to the form usually quoted:
In the picture of nearest neighbors behind the model,
W must be square and symmetric, which a validating inner constructor enforces; the reason it is inner rather than outer is recorded in the source, and it is a mistake worth not repeating.
The threshold is
For a symmetric binary, the molar Gibbs energy of mixing is
which at
This matters in practice because nothing in the activity expression detects it. Past the threshold it goes on returning values, and those values describe a single phase that should have split. The check is therefore made at construction: SolidSolutionPhase evaluates the second derivative and refuses a model that unmixes at the temperature given, naming the interval, unless the phase is declared with two instances (section 6) or the check is switched off with check_convexity = false.
4. RedlichKisterModel — a binary that need not be symmetric
A regular solution is symmetric by construction: swapping the end-members leaves
with
RedlichKisterModel requires exactly two end-members, and SolidSolutionPhase refuses the combination at construction rather than letting a ternary reach an expression written for a binary. For three or more end-members the choices are the ideal model, RegularSolutionModel with a full
Published dimensionless parameters, and the temperature
Cemdata18 gives its non-ideal AFt and AFm binaries dimensionless Guggenheim parameters data/literature/Lothenbach2019.json), and no temperature dependence for them. The package turns them into
using ChemistryLab, DynamicQuantities, Printf
p = literature_row("Lothenbach2019", "guggenheim_parameters", "AFm SO4/OH")
α0, α1 = ustrip(p.alpha0), ustrip(p.alpha1)
rk(T) = RedlichKisterModel(a0 = α0 * R_GAS * T, a1 = α1 * R_GAS * T)
@printf("printed gap: %.2f to %.2f\n", ustrip(p.gap_from), ustrip(p.gap_to))
println(" °C   a in J/mol held (the package)   α held")
for T_C in (5, 25, 50, 80)
    T = T_C + 273.15
    held_a = common_tangent(rk(298.15); T)
    held_α = common_tangent(rk(T); T)
    @printf("%3d       %.3f to %.3f            %.3f to %.3f\n", T_C, held_a..., held_α...)
end
printed gap: 0.50 to 0.97
°C a in J/mol held (the package) α held
5 0.478 to 0.977 0.500 to 0.970
25 0.500 to 0.970 0.500 to 0.970
50 0.528 to 0.959 0.500 to 0.970
80 0.565 to 0.943 0.500 to 0.970Both give the printed gap at 25 °C. The package's choice narrows it as the temperature rises, from 0.48–0.98 at 5 °C to 0.57–0.94 at 80 °C (the mole fraction of C₄AH₁₃, the OH end, as the edges are printed); the other keeps it where Cemdata18 prints it.
5. What the three models do to an activity
Evaluated over the whole composition range in Solid solution models, in numbers, with
A positive
The ideal case is the one to keep in mind for cement. An end-member at
6. The spinodal, the common tangent, and why one amount is not enough
Sections 3 and 4 gave models whose excess term can be strong enough to make the molar Gibbs energy of mixing concave over an interval. This section is about what that means for the equilibrium, because it is not a detail of the model: it changes what the answer is.
Convexity is the hypothesis under everything else
For a binary write the molar Gibbs energy of mixing, in units of
The ideal part is convex everywhere — its second derivative is
the phase is inside its spinodal, and there the straight line joining two compositions lies below the curve between them. A system at an intermediate overall composition therefore lowers its energy by separating into those two, and the minimum of
For the symmetric regular solution of section 3 the criterion is exactly spinodal_interval evaluates the second derivative on a grid rather than in closed form, which is why the same routine covers the asymmetric Redlich-Kister case without a separate derivation.
The answer inside a gap is the common tangent
Which pair? Write
the conservation of the two end-members. With a multiplier
The four equations say one thing: the straight line common_tangent solves:
The multipliers are chemical potentials. The tangent to
Nothing requires the two points to be at the same height:
What the split gains follows from the lever rule,
The common tangent construction is wider than the spinodal: the binodal
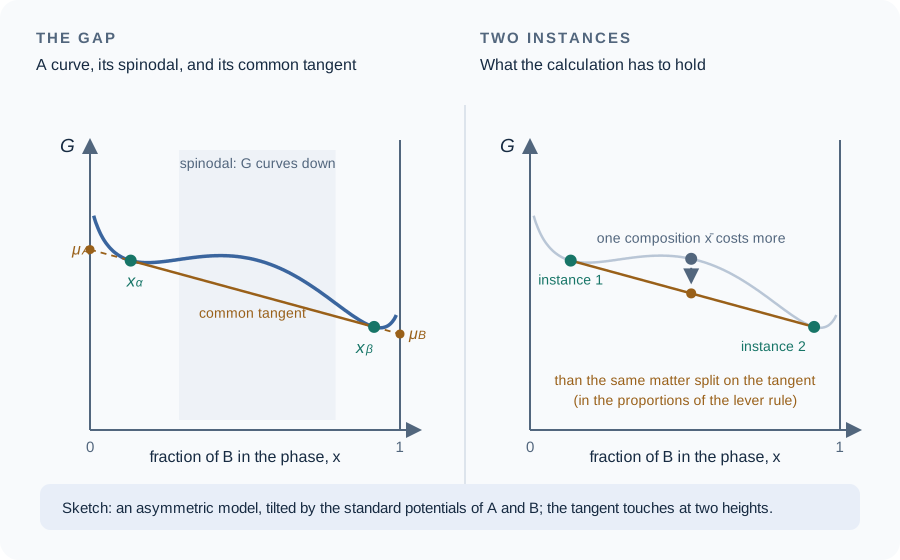
The right panel states the difficulty the rest of this section resolves: inside the gap the equilibrium holds the phase at two compositions at once, in the proportions of the lever rule, which a formulation carrying one amount per end-member has no means to express.
Computing the pair from the model alone
The pair is not found by minimizing. It is computed from the model alone, by common_tangent: two equations in two unknowns, solved by Newton with ForwardDiff supplying the derivatives. Nothing about the rest of the chemical system enters — not the aqueous solution, not the other phases, not the element budget — which is why it costs microseconds and can be used as the starting point of a full equilibrium rather than as its result.
The construction is that of Glynn and Reardon (1990) for binary solid solutions, which PHREEQC (Parkhurst and Appelo, 2013) uses as well. It was validated here against a case with a closed form: for a symmetric model the pair must be symmetric about
miscibility_split then applies the lever rule. Inside the gap the two compositions are fixed and only their proportions move with the overall composition — which is what makes a miscibility gap flat in a phase diagram, and what a single-composition answer cannot reproduce at any resolution.
Why a formulation with one amount per species cannot hold it
The composition vector of a ChemicalSystem has one entry per species, so a species belongs to exactly one phase and a phase has exactly one composition. A common-tangent pair is two compositions of one substance, and there is no way to write that down with one amount each.
So the formulation has to be given the substance twice. That is what SolidSolutionPhase(...; instances = 2) does: ChemicalSystem builds a second copy of each end-member under a derived symbol (monosulphate12#2) sharing the same thermodynamic record, and each copy belongs to its own phase. The groups stay disjoint, the duplicated columns of the conservation matrix are copies of columns already there — so the row rank, and with it conservation, is untouched — and the minimization is free to populate either lobe or both.
It is refused for a convex model, and the reason is worth stating because it is not conservatism. Two instances of a convex phase are degenerate: the energy is the same however the amount is split between them, so the minimum becomes a flat manifold and the optimizer is asked to pick a point on it arbitrarily. Inside a spinodal the common-tangent pair is unique and no such direction exists.
Michelsen's tangent-plane distance, which is what detects it
Representation is one problem; knowing that it is needed is another. A phase sitting inside its own spinodal satisfies every first-order condition — its end-members are stationary, the element balance closes, nothing absent is supersaturated — so stationarity alone certifies a non-minimum.
The test that separates them is Michelsen's (Michelsen, 1982): for a trial composition
measures how far the Gibbs surface at
OptimaSolver's phase_split_measure computes this for every present mole-fraction phase and folds the result into the certificate. Concretely: check_convexity = false stops being a silent loss of the proof — a concave declaration that does unmix now fails to certify, with the phase named.
A second composition, declared or added when needed
The certificate's measure of a phase is the sum over its members of their activities from the dual solution divided by their activity coefficients from the primal one, an estimate of the mole fraction the phase would take:
It is the phase stability index phase_split_measure.
Holding a gap takes the phase twice. Cemdata18 ships the AFm and AFt binaries under two names each, so that the database itself declares each binary twice; instances = 2 declares the second composition with a keyword. instances = :auto asks for less: the phase is solved with one composition, and a second instance is added only when the certificate finds it wanting to split. On a binary whose overall composition the element budget holds inside the gap, calcite and magnesite in equal amounts, that finds the common-tangent pair:
using ChemistryLab, DynamicQuantities, Printf
sp = Dict(symbol(s) => s for s in build_species(datapath("slop98-inorganic-thermofun.json"); verbose = false))
gap = RedlichKisterModel(a0 = 14_000.0)
carbonate_ss = SolidSolutionPhase("carbonate", [sp["Cal"], sp["Mgs"]]; model = gap, instances = :auto)
cs = ChemicalSystem([sp[s] for s in split("H2O@ H+ OH- CO2@ HCO3- CO3-2 Ca+2 Mg+2 Cal Mgs")],
                    ["H2O@", "H+", "Ca+2", "Mg+2", "CO3-2", "Zz"]; solid_solutions = [carbonate_ss])
st = ChemicalState(cs)
set_quantity!(st, "H2O@", 1.0u"kg"); set_quantity!(st, "Cal", 0.025u"mol"); set_quantity!(st, "Mgs", 0.025u"mol")
eq, cert = equilibrate_certified(st; b = budget(st))
n = ustrip.(us"mol", eq.n)
@printf("certified %s, instances added to %s\n", cert.optimal, cert.instances_added)
for (g, ph) in zip(eq.system.ss_groups, eq.system.solid_solutions)
    @printf("  %-12s %.5f mol, x(calcite) = %.4f\n", name(ph), sum(n[g]), n[g[1]] / sum(n[g]))
end
@printf("common tangent of the model: %.4f and %.4f\n", common_tangent(gap)...)
certified true, instances added to ["carbonate"]
carbonate 0.02495 mol, x(calcite) = 0.9963
carbonate#2 0.02486 mol, x(calcite) = 0.0037
common tangent of the model: 0.0037 and 0.9963It does not make a nonconvex problem easy: the certificate names a phase to split only once the one-composition solve has converged.
The executed counterpart of this section is the miscibility-gap page, which runs one cement three ways and reports the certificate each time.
7. Mixing on several sites: SublatticeModel
Sections 1 to 4 mix end-members. In a C-S-H the entities that actually mix are smaller: a few structural positions of the silicate chain and of the interlayer, each held by one species or another. An end-member is then one particular filling of those positions, and mixing is random on each position separately. That is the model of Kulik (2011) for CSH3T and of Myers et al. (2014) for the CNASH gel, and it is what SublatticeModel implements.
The model. A formula unit has sites
The implication is excess_ln_gamma_expression returns for this model.
CSH3T. Two bridging-tetrahedral sites hold either Si or Ca. TobH puts Si on both, T2C puts Ca on both, and the ordered T5C puts Ca on the first and Si on the second (Kulik, 2011; Eq. 19). The Cemdata18 records are half of its formula units, so on them each site counts one half:
using ChemistryLab
subs = Dict(symbol(s) => s for s in build_species(datapath("cemdata18-thermofun.json"); verbose = false))
csh3t = sublattice_model("Kulik2011:csh3t", [subs[n] for n in ("CSH3T-TobH", "CSH3T-T5C", "CSH3T-T2C")])
csh3t.multiplicity
2-element Vector{Float64}:
0.5
0.5sublattice_model finds that factor itself, by comparing each record's formula with the one Kulik (2011) prints, and refuses a record that is not such a multiple. At
site_fractions(csh3t, [0.2, 0.5, 0.3])2-element Vector{Vector{Float64}}:
[0.2, 0.8]
[0.7, 0.3]and the fictive activity coefficient of T5C is Eq. (20) of Kulik (2011), halved:
The block below builds that expression from the model and subtracts the formula above; the difference simplifies to zero:
using Symbolics
x = [Symbolics.variable(:x, i) for i in 1:3]   # x₁ = TobH, x₂ = T5C, x₃ = T2C
λ = excess_ln_gamma_expression(csh3t, 2, 3)
simplify(expand(λ - (0.5log(x[1] + x[2]) + 0.5log(x[3] + x[2]) - log(x[2]))))
Two properties the solver relies on.
Convexity.
is a sum of convex functions of the site fractions, which are linear in , so it is convex, and strictly so when the occupancy matrix (one row per site and species, one column per end-member) has full column rank, stored as rank. The activities are the gradient of that energy, so a certificate keeps its:global_minimumscope.The dilute limit. As
alone carries vanish with it: the multiplicity of those sites, stored as exponents. T5C owns no species of its own,
csh3t.exponents3-element Vector{Float64}:
0.5
0.0
0.5What the solver does with it. The composition of a mixing phase is found by successive substitution, and on a sublattice model that iteration diverges: its rate is set by the sum of the multiplicities, nine for the CNASH gel. Such a phase is therefore inverted by Newton's method, and the members with SolutionPhase, keywords newton and bounded_members). A system with a sublattice phase is first solved with that phase mixing its end-members ideally, and the answer is the start of the sublattice solve.
data/solid_solutions.toml ships CNASH_ss as the model of Myers et al. (2014) and CSH3T as Cemdata18 ships it, ideal between end-members; sublattice_model("Kulik2011:csh3t", members) gives the site form.
8. Sites with the energies of the compounds: CompoundEnergyModel
Section 7 mixes ideally on the sites and keeps the end-members' own standard energies,
leaves every site fraction unchanged, yet its standard Gibbs energy is not zero. The compound energy formalism, in which Kulik et al. (2022) write the model, gives such a reaction its energy. CompoundEnergyModel implements it.
The Gibbs energy. The end-members must be every compound of the sites, each once. Compound
The first term, the reference surface, equals
The activities. With
On the reference surface
The middle term vanishes at every compound. It also does not change when each
Three consequences.
The amounts are not unique. Every split of the same site fractions between the members has the same
and the same element content: in CASH+, six amounts for three independent site fractions and one total. Left to choose, the solver returns one of them, which one depending on where the search started. It is therefore given one split, the product of the site fractions, is the divergence of from the product of its own site fractions: it is never negative, and it vanishes only at that product, where its gradient , at the same cost; what adds is an answer that does not depend on the start. Convexity must be decided. The reference surface is multilinear, and a reciprocal energy can make
concave somewhere. In the site fractions, each site contributes a curvature of at least of its interactions , and with two sites the reference surface couples them through a constant matrix , the of the compounds projected on the two tangent spaces. So is convex when :global_minimumscope as far as the gel goes; the bound does not decide CASH+NK, whose sampled Hessian shows no concave point, and its scope is then:local_minimumor:kkt_point, according to the curvature at the answer (mixing_convexity).Euler's relation holds.
, as it must for a Gibbs energy.
The block below checks the gradient and the last point on the core model, at a composition of the six members. The activities are compared with the gradient of
using ChemistryLab, DynamicQuantities, ForwardDiff, LinearAlgebra
subs = Dict(symbol(s) => s for s in build_species(datapath("cemdata18-cashplus.json"); verbose = false))
members = ["TSvh", "TSCh", "Tvvh", "TCvh", "TvCh", "TCCh"]
cash = compound_energy_model("Kulik2022:cashplus", [subs[m] for m in members])
T = 298.15
g = [ustrip(us"J/mol", subs[m][:ΔₐG⁰](T = T * u"K", P = 1.0e5u"Pa"; unit = true)) for m in members] ./ (ChemistryLab.R_GAS * T)
o = cash.lattice.occupancy
function nG(n; split = true)
    x = n ./ sum(n)
    y = site_fractions(cash, x)
    G = sum(g[j] * y[1][o[1, j]] * y[2][o[2, j]] for j in 1:6) + sum(v * log(v) for ys in y for v in ys)
    G += sum(W / (ChemistryLab.R_GAS * T) * y[s][i] * y[s][l] for (s, i, l, W) in cash.interactions)
    split && (G += sum(v * log(v) for v in x) - sum(v * log(v) for ys in y for v in ys))   # D(x)
    return sum(n) * G
end
n = [0.10, 0.25, 0.05, 0.20, 0.15, 0.25]
lna = ChemistryLab._ss_log_activities!(zeros(6), 1:6, n ./ sum(n), cash, T, 0.0, g)
μ = ForwardDiff.gradient(nG, n)
y = site_fractions(cash, n ./ sum(n))
xp = [y[1][o[1, j]] * y[2][o[2, j]] for j in 1:6]          # the product split
(gradient = maximum(abs, lna .+ g .- μ), euler = dot(n, μ) - nG(n),
 D_at_product = nG(xp) - nG(xp; split = false))
(gradient = 6.821210263296962e-13, euler = 0.0, D_at_product = 0.0)All three are at the rounding of the arithmetic. The model and its data are in data/literature/Kulik2022.json, the sodium and potassium of Miron et al. (2022), Miron et al. (2022) in Miron2022a.json and Miron2022b.json, and the twelve end-members in the database cemdata18-cashplus.json. The CASH+ page computes the C-S-H in water and in alkali solutions with it.
9. More than two end-members
Asymmetric mixing of any number of end-members: SubregularSolutionModel
The regular model of section 3 is symmetric on every edge, and the Redlich–Kister model of section 4 is asymmetric but binary. Helffrich and Wood (1989) write the asymmetric Margules model for any number of end-members (their Eq. 5′):
On the edge
Helffrich and Wood (1989) give the activity coefficients (their Eq. 6′) without the intermediate steps. They are short once the excess is written on the simplex, where
so each pair is a Redlich–Kister term of the first order,
The formula holds whatever
and a triple,
with _excess_ln_gamma(::SubregularSolutionModel, …) term by term. Setting
A series for each pair: MulticomponentRedlichKisterModel
Redlich and Kister (1948) write the excess of a binary as a series in the difference of the two mole fractions, which changes sign when the components are exchanged, and that of a ternary as the sum of its three binaries, each evaluated at the mole fractions of the ternary as they are, plus a term in the product of the three (their Eqs. 17 and 18):
The order of a pair matters for its odd terms, which change sign with it, and the order of a triple for
The activity coefficients follow from Eq. (14) of Redlich and Kister (1948), the partial molar derivative written in mole fractions, which is the formula of the subregular model above. For a pair, with
and for a triple, with
_excess_ln_gamma(::MulticomponentRedlichKisterModel, …). Two end-members give the model of section 4 with
Redlich and Kister (1948) also give the ratio of two activity coefficients in a ternary (their Eq. 19). Its term in
Unequal sizes: VanLaarModel
Holland and Powell (2003) make the regular model asymmetric by giving each end-member a size
Only the ratios of the sizes matter. Equal sizes give back the regular model of section 3, which Holland and Powell (2003) call the symmetric formalism, and two end-members give the van Laar binary, asymmetric as soon as the sizes differ. The papers that fit
Holland and Powell (2003) write the activity coefficients (their Eq. 2) with interactions rescaled by the size of the end-member considered,
a quadratic form over a linear one. The excess of
which is _excess_ln_gamma(::VanLaarModel, …). With equal sizes
Neither this model nor the series above has the Redlich-Kister form of section 4 with three coefficients, so their binary is scanned for a spinodal and its common tangent found from the excess itself,
Convexity with more than two end-members
A one-dimensional scan decides a binary. With mixing_convexity answers it by model.
For a regular model,
:undecided. The subregular, the Redlich-Kister and the van Laar models are decided by that search alone beyond two end-members: their Hessian depends on the composition through the asymmetric and the ternary terms, and no bound of the kind above is written for them.
using ChemistryLab, DynamicQuantities, Printf
RT = ChemistryLab.R_GAS * 298.15
W(a, b, c) = [0.0 a b; a 0.0 c; b c 0.0] .* RT
[mixing_convexity(RegularSolutionModel(W(1.9, 1.9, 1.9)), 3).verdict,
 mixing_convexity(RegularSolutionModel(W(3.0, 0.0, 0.0)), 3).verdict]
2-element Vector{Symbol}:
:convex
:nonconvexA phase whose verdict is :nonconvex is refused at construction, as a concave binary is, and admitted with two instances. The certificate of an answer is not scoped :global_minimum unless every mixing phase is proved convex; it is then :local_minimum where the Hessian over the directions that conserve matter is positive definite at the answer, and :kkt_point otherwise.
A member that is a mixture of two others
Two different things can make one end-member the mixture of two others in composition. In an ordered member the difference of Gibbs energy is the point: the pentameric T5C of CSH3T is the average of TobH and T2C less an ordering energy (Kulik, 2011; Eq. 18), and the siliceous hydrogarnet C3AS0.41H5.18 lies between C3AH6 and C3AS0.84H4.32. When the Gibbs energy is the average as well, the phase holds one substance twice, once as a member and once as a mixture of two, and ideal mixing counts its configurations twice. SolidSolutionPhase warns when that holds within
subs = Dict(symbol(s) => s for s in build_species(datapath("cemdata18-thermofun.json"); verbose = false))
g(s) = ustrip(us"kJ/mol", subs[s][:ΔₐG⁰](T = 298.15u"K", P = 1.0e5u"Pa"; unit = true))
@printf("MgAl-OH-LDH  M6A - (M4A + M8A)/2  = %+.4f kJ/mol\n",
        g("M6A-OH-LDH") - (g("M4A-OH-LDH") + g("M8A-OH-LDH")) / 2)
@printf("CSH3T        T5C - (TobH + T2C)/2 = %+.2f kJ/mol\n",
        g("CSH3T-T5C") - (g("CSH3T-TobH") + g("CSH3T-T2C")) / 2)
MgAl-OH-LDH M6A - (M4A + M8A)/2 = +0.0003 kJ/mol
CSH3T T5C - (TobH + T2C)/2 = -4.35 kJ/molThe LDH difference, 0.3 J/mol, is about 10⁻⁴ RT, where the −4.35 kJ/mol of T5C is an ordering energy. The shipped LDH entry keeps the published model with acknowledge_degenerate = true.
10. How a solid solution is declared
A SolidSolutionPhase names its end-members and carries a model:
phase = SolidSolutionPhase("CSHQ", [sp["CSHQ-JenD"], sp["CSHQ-JenH"],
                                    sp["CSHQ-TobD"], sp["CSHQ-TobH"]];
                           model = IdealSolidSolutionModel())
Every end-member must be AS_CRYSTAL, and the phase is checked against its model's arity and against the convexity of its mixing energy at construction; instances is how a declaration asks for more than one coexisting composition, which section 6 is about. Sets of end-members with their interaction parameters are read from a TOML through build_solid_solutions, which is also the only interaction-parameter file format in the package — data/solid_solutions.toml is the shipped example.
Two consequences to keep in mind when reading results:
a
ChemicalSystemgroups end-members intoss_groups, and every aqueous activity model must call the solid-solution branch or the end-members come back withan end-member's
saturation_indicesentry is relative to its current mole fraction, since its activity is. For an end-member sitting at the solver's lower bound that is a statement about a vanishing phase, not about whether the solid solution would form.
See also
Thermochemistry — where
enters Activity models — the aqueous half
Solving an equilibrium — declaring and solving with solid solutions